LNP Manufacturing Market: Growth, CDMO Trends and What Developers Need to Know

LNP manufacturing has become one of the fastest-growing areas in modern pharmaceutical development. Lipid nanoparticles (LNPs) have moved from a niche formulation approach to one of the most validated platforms for delivering complex therapeutic payloads, and demand for specialized development and manufacturing partners has grown alongside them.

According to market research firm InsightAce Analytic, the global lipid nanoparticles (LNP) CDMO market was valued at USD 205.4 million in 2024 and is projected to reach USD 803.6 million by 2034, growing at a CAGR of 14.8%. 

The firm attributes this expansion to rising demand for lipid-based drug delivery, increasing investment from key players, and the growing complexity of the therapies these systems are built to carry, as set out in its market report, for which a free brochure is also available to download.

The state of LNP manufacturing

The appeal of lipid-based delivery is well established. Since the first clinical approval of a liposomal product in the 1990s, lipid nanoparticles have proven their ability to encapsulate a wide range of therapeutic agents, from small molecules and peptides to nucleic acid therapies and antibodies. 

Their biocompatibility, feasibility of scale-up, and clinical track record have made them one of the most trusted platforms in drug delivery.

The growth the report describes is driven less by novelty and more by outsourcing. Pharmaceutical and biotech developers are increasingly trusting formulation development, process optimization, analytical testing, and GMP production to specialized partners rather than building that capability in-house. 

At the same time, the therapies themselves keep getting more complex, which raises the bar for encapsulation efficiency, particle characterization, and quality control. That combination is what sustains steady demand for CDMOs with genuine lipid chemistry expertise.

Scale changes everything

The InsightAce report segments the market by scale of operation: preclinical, clinical, and commercial. In practice, and reading beyond the report’s segmentation, what a sponsor needs from a partner changes significantly across those stages, from formulation feasibility and early process development to GMP clinical supply and, ultimately, commercial-scale manufacturing.

At the preclinical stage, the priority is feasibility and formulation design. This is where lipid selection, particle engineering, and early encapsulation work happen, and where a sponsor benefits most from a partner with hands-on experience across different lipid chemistries. Speed and flexibility matter more than volume, and decisions made here shape everything downstream.

At the clinical stage, the emphasis shifts to reproducibility, analytical rigor, and GMP production. The formulation that worked at the bench now has to be manufactured to a clinical standard, characterized against the critical quality attributes regulators expect, and documented for submission. A sponsor needs a partner whose process development was built with scale-up in mind from day one, so nothing is lost in the transfer from feasibility to GMP clinical supply.

At the commercial stage, the questions change again: sustained capacity, supply chain resilience, and large-volume consistency. This is a distinct challenge with its own infrastructure demands, and it is worth being clear-eyed about which partners are positioned for development and clinical supply versus full commercial output.

For a development-focused program, the practical takeaway is to match the partner to the stage. Ardena, for example, concentrates on the earlier part of that journey, taking lipid-based nanoparticle programs from first formulation through to GMP clinical supply, with process development designed for scale-up from the outset.

What the report calls hard

The same report is candid about what constrains this market, and it is worth restating those points plainly rather than glossing over them. InsightAce identifies the high cost of outsourced services, a lack of necessary infrastructure, the need for specialist expertise in nanoparticle production, and the complexity of the manufacturing procedures themselves as the main factors holding growth back.

None of these are trivial. Lipid nanoparticle manufacturing sits at the intersection of formulation science, analytical chemistry, and sterile production. 

Bringing those together within a single development pathway requires specialist infrastructure and expertise. Encapsulation has to be efficient and reproducible. 

Characterization has to satisfy regulators as well as scientists. And the whole process has to hold together under GMP conditions. These are exactly the areas where inexperience becomes expensive, which is why the choice of partner carries so much weight.

The provider landscape

In its competitive landscape, InsightAce identifies Ardena among the major key players in the global LNP CDMO market. InsightAce subsequently highlighted Ardena in a LinkedIn overview of ten LNP CDMO companies, further recognizing its presence within the specialist provider landscape.

Ardena’s place in that landscape reflects its focus rather than its size. As an integrated CDMO and bioanalytical CRO, it develops lipid-based nanoparticles including LNPs, liposomes, and lipid micelles, with process development that spans phospholipids, neutral lipids, cationic lipids, and hybrid lipid systems. Its analytical scientists characterize particles against the full set of critical quality attributes, from particle size distribution and zeta potential to lipid quantification and confirmation of encapsulated payload, and its dedicated nanomedicine platform operates from a purpose-built GMP facility designed to meet current regulatory requirements, including Annex 1. 

The consistent thread is a focus on technically demanding molecules and formulations that require specialist development, analytical and manufacturing expertise.

What to ask a potential partner

Because scale and difficulty are the two forces that shape this market, they also make a good starting point for evaluating a partner. A few questions that follow directly from the points above:

Which stage are you built for?
Ask a prospective partner to be specific about where they operate, from preclinical feasibility through clinical GMP supply, and be wary of vendors who claim to do everything equally well.

Do formulation, analytics, and GMP production sit under one roof?
Handoffs between separate providers are where momentum and know-how get lost, so integrated capability is worth a lot.

How is scale-up handled?
A partner should be able to explain how a bench formulation moves to GMP without being fundamentally reworked, because scale-up is one of the most technically sensitive points in LNP development.

How deep is the lipid chemistry experience?
Given the specialist expertise the report identifies as a constraint, a track record across different lipid systems and payloads is a meaningful differentiator.

Looking ahead

With the market projected to more than triple in value over the next decade, LNP manufacturing will remain a strategically important capability for the industry. The developers who move fastest will be those who pair a strong therapeutic idea with a partner matched to their stage, one that already knows how to formulate, characterize, and manufacture lipid nanoparticles to a clinical standard.

If you are developing a lipid-based program, Ardena’s scientists work on exactly this kind of formulation and manufacturing challenge.

Market data referenced from InsightAce Analytic, “Lipid Nanoparticles (LNPs) CDMO Market Size, Share and Growth Analysis 2025 to 2034.” See the full report and free brochure and the publisher’s LinkedIn overview.
How to Evaluate HPAPI Contract Manufacturing Companies in 2026: A Technical and Risk-Based Guide

HPAPI contract manufacturing companies are not interchangeable, and choosing the right one is among the highest-stakes decisions in a potent-compound program. 

Pick the wrong partner and it surfaces later: as a containment finding, a failed cleaning validation, or a technology transfer that stalls before your first batch. 

This guide is about how to make that choice well, covering the criteria, trade-offs and questions that separate a genuinely capable partner from a persuasive sales deck. It is deliberately not a description of what any one provider makes.

Start with containment you can verify, not containment that is claimed

Potency is classified using occupational exposure banding, the industry framework that sorts compounds by how hazardous they are to handle. It runs from band 1, the least hazardous, to band 5, the most potent, and some classification systems add a sixth band for ultra-potent conjugates. 

Band 4 covers compounds with an occupational exposure limit between roughly 1 and 10 micrograms per cubic meter of air, and band 5 covers those below 1 microgram, the point at which standard suites can no longer protect operators or reliably prevent cross-contamination. 

From band 4 upward, containment, cleaning and trace-level analysis all have to be purpose-built. The band your molecule sits in dictates the engineering a partner actually needs: sealed isolators and closed transfers, not respirators and written procedures. 

Ask every shortlisted provider for documented surrogate containment test data at your band, generated at the specific site that would run your program rather than a corporate average. 

A clean inspection record and containment evidence at the exact facility that will make your batches matter more than a global footprint. 

A partner that treats containment designed in from the start as an engineering discipline is far more defensible, to regulators and to your pharmacovigilance team, than one that leans on protective equipment.

Match the partner to your molecule and its stage

Potency rarely travels alone. Many potent compounds are also poorly soluble, structurally complex, or destined to become a conjugate, and each raises a different question. 

For a difficult route, ask how a provider approaches the efficient synthesis of highly potent compounds at contained scale, and whether one team can carry the chemistry from milligrams to kilograms without a mid-program site change. If your molecule is a payload or linker, generic potent-handling experience is not enough, so look for dedicated ADC and XDC programs. 

If you need drug substance, probe the depth of their HPAPI synthesis record for compounds like yours. Start this conversation before the chemistry is locked; early input from a partner with high potency and controlled substances experience can shape a route that is easier and safer to contain later.

Treat cleaning validation as a patient-safety question

Where equipment is shared, cleaning validation is what protects the next patient from your compound. 

Confirm that acceptance limits come from compound-specific health-based exposure limits set by a qualified toxicologist, not legacy parts-per-million rules, and that the analytical method can reliably detect residues below the calculated carryover. 

Then ask whether your compound would run on dedicated or campaign equipment, and what that means for scheduling and cross-contamination risk.

Look beyond the suite: supply chain and change control

Two factors that buyers routinely underweight sit outside the containment suite. The first is starting-material and supply-chain transparency. 

For compounds built through complex multi-step synthesis or controlled precursors, a partner’s ability to source, qualify and document those materials affects both regulatory risk and continuity of supply, and reshoring pressure has turned provenance into a live question rather than a formality. 

The second is change control. A manufacturing change at your partner’s site can trigger a regulatory variation for your own product, so ask how changes are assessed, communicated and timed, and whether the site’s change history has ever caused a quality problem.

Questions that separate capability from a sales deck

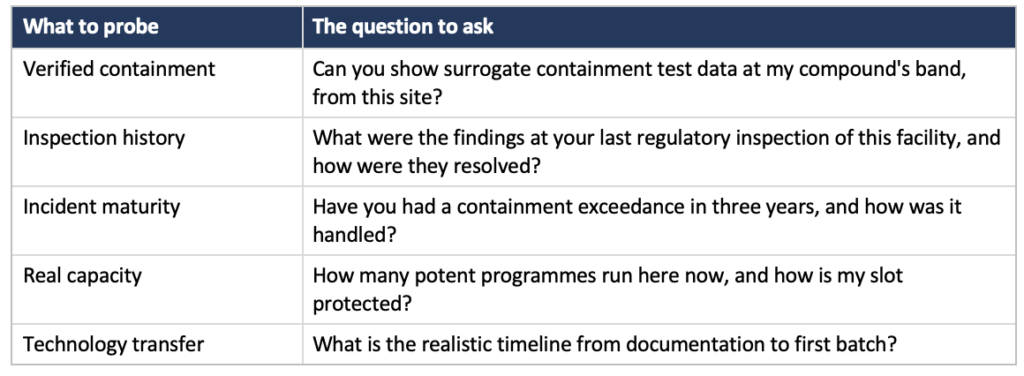
When you compare HPAPI contract manufacturing companies side by side, the differences that matter rarely appear on the capability slide. These do:

hpapi contract manufacturing companies

The questions buyers most often skip are the human ones: who your project manager is, what happens when they leave, and how the site resolves competing priorities when a larger client wants the same suite the same week. For the quality-system and project-management ground that applies to any first manufacturing campaign, our due diligence checklist for Phase I CDMO selection is a useful companion to this guide.

The bottom line

The right partner is the one whose evidence matches your molecule’s band, stage and complexity, and who shows you the data before you have to ask twice. Ardena publishes that evidence for its own high potency site: occupational exposure measurements with a model compound, four segregated suites, and a facility that has passed every regulatory inspection since 2012.

Related reading

Encapsulating Challenging APIs: High Potency and Low Solubility

ADC and XDC Drug Development: What CDMOs Need to Deliver

Understanding Highly Potent APIs and Safe Handling

The Virtual Biotech Model: How to Build a Drug Program Without a Lab

The Model That Changed Biotech

Twenty years ago, a credible pharmaceutical development program needed laboratories, equipment, and scientists on the payroll. The idea that a five-person team could advance a drug candidate from lead optimization to Phase II without owning a single centrifuge would have seemed implausible.

Today it is routine. The virtual biotech model, built on outsourcing relationships with CDMOs, CROs, and specialist consultants, has become one of the dominant organizational forms in early-stage pharmaceutical development. It allows small teams to access world-class scientific and manufacturing capabilities without the capital overhead, and it lets investors fund science rather than infrastructure.

It also fails regularly, for predictable reasons that have nothing to do with the science.

What Goes Wrong in Virtual Biotech Programs

Too Many Vendors, Not Enough Oversight

The instinct of many virtual biotech teams is to find the best vendor for each individual piece of work. Best formulation house for development, best CRO for animal studies, best CMO for manufacturing. The result is a program spread across five or six organizations with no single point of accountability and a coordination overhead that consumes the management team.

No Internal CMC Expertise

A virtual biotech can outsource the execution of CMC work. It cannot outsource the judgment about what CMC work is needed. Without someone on the team who understands what a regulator will expect to see in Module 3, the program can spend months generating data that does not answer the right questions.

The Tech Transfer That Should Not Have Existed

Many virtual biotechs develop a molecule with one vendor, then transfer it to a different GMP manufacturing partner because the first vendor does not have GMP capability. Every tech transfer is a risk event. Data is interpreted by new people, processes are reproduced on different equipment, and unexpected results emerge. The transfer that could have been avoided by choosing an integrated partner from the start costs months and money to resolve.

What Successful Virtual Biotechs Do Differently

Success FactorWhat It Looks Like in Practice
Choose an integrated partner earlySelect a CDMO with development and GMP manufacturing capability at the start; avoid planned tech transfers between development and manufacturing
Keep CMC expertise internalHire or retain a Head of CMC or CMC consultant with regulatory filing experience; do not rely on the CDMO to own regulatory strategy
Define deliverables, not activitiesContract for outcomes (Phase I CMC package) rather than activities (12 months of stability work); gives the CDMO flexibility to design the most efficient program
Build in formal decision gatesSchedule quarterly scientific reviews where the program strategy is assessed against the data, not just the timeline
Use project management tools the CDMO actually supportsDo not assume the CDMO will use your project management system; agree on reporting format and frequency before work starts
Plan for the unexpectedBuild contingency time and budget into the plan; virtual programs with no slack are brittle when something goes wrong

The Integrated CDMO as a Virtual Biotech’s Scientific Department

For a five-person virtual biotech, the CDMO is not just a service provider. It is, functionally, the scientific organization. The quality of the science that goes into the program, the data generated, the decisions made, the regulatory strategy pursued, is largely determined by the quality and engagement of the CDMO team.

This is why the consultative tone that characterizes good CDMO partnerships matters more for virtual biotechs than for large pharma. A CDMO that executes instructions without pushing back when the science suggests a different approach is not providing the partnership a virtual biotech needs. A CDMO that raises the question about polymorph stability before the first GMP batch is planned, that flags the dissolution method issue before the IND is filed, is providing something genuinely valuable.

How Ardena Supports Virtual Biotech Programs

Ardena’s model is particularly well suited to virtual biotech programs. The integrated multi-site network covers solid state research, formulation development, analytical services, GMP manufacturing, bioanalysis, and regulatory support under a single project management framework. For a virtual biotech that does not have scientific departments of its own, Ardena provides the depth of capability that would otherwise require five separate vendor relationships.

Quality Agreements in Pharmaceutical Outsourcing: What They Must Cover

The Document Nobody Reads Until Something Goes Wrong

Quality agreements are one of those documents that get negotiated carefully, signed by the right people, filed away, and never looked at again. Until something goes wrong.

When a GMP deviation occurs, when a batch fails, when a regulatory inspection raises a question about oversight, the quality agreement is the document that defines who was responsible for what. If it is vague, incomplete, or has not been reviewed since the program started, the consequences are predictable.

A good quality agreement is not just a legal formality. It is a genuine operational tool that prevents disputes, clarifies accountability, and ensures that GMP responsibilities are distributed logically between the sponsor and the CDMO.

The Regulatory Requirement

EU GMP Chapter 7 on outsourced activities and the FDA’s guidance on contract manufacturing both require written quality agreements between sponsors and their contract manufacturers. The agreement must define the GMP responsibilities of each party clearly and must be reviewed and approved by the quality units of both organizations. It is not sufficient for a general commercial contract to contain quality provisions; a standalone technical or quality agreement is the regulatory expectation.

What a Strong Quality Agreement Covers

SectionWhat It Should DefineCommon Weakness
Scope of workExactly which activities are covered; what is in scope and out of scope for the agreementScope defined too broadly; changes in the program not captured by agreement updates
GMP responsibilitiesWhich party is responsible for each GMP activity: batch record review, release, deviation investigation, CAPAJoint responsibilities listed without specifying the primary responsible party
Change controlWhich changes require sponsor notification; which require sponsor approval before implementationCDMO retains right to make process changes without sponsor approval; sponsor not notified in time
Deviations and OOSTimelines for notification; who investigates; who approves the investigation reportNotification timelines too loose; sponsor does not receive deviation reports until batch review
Regulatory inspectionsWhat each party does when the other is inspected; how the agreement is referenced in submissionsNo provision for inspection support; sponsor not informed of CDMO inspection findings
StabilityWho owns the stability program; who has responsibility for out-of-trend resultsStability ownership split between parties without clear escalation path
Batch releaseAuthorized person responsibilities; what happens when the sponsor and CDMO disagree on releaseRelease responsibility assumed but not explicitly assigned; AP not named
Annual product reviewWhich party leads the APR; what data the CDMO provides; timelinesAPR responsibility not assigned; CDMO data not available when sponsor needs it

The Change Control Problem

Change control is the section of quality agreements that generates the most disputes in practice. The question is simple: when the CDMO wants to change something about the manufacturing process, the equipment, the supplier, or the facility, what does it need to tell the sponsor, and when?

From the CDMO’s perspective, the ability to make routine operational improvements without seeking sponsor approval for every change is a reasonable operational requirement. From the sponsor’s perspective, any change to the process that manufactured their clinical batches is a potential CMC filing issue that could trigger a regulatory amendment.

The resolution is a tiered change control classification. Minor changes are notified; significant changes require sponsor review; major changes require sponsor approval before implementation. What counts as minor, significant, or major should be defined explicitly in the agreement, not left to interpretation.

Reviewing and Updating the Agreement

Quality agreements should be reviewed at least annually and updated whenever the scope of work changes materially. A quality agreement written at the start of a Phase I program will not adequately cover the same program at Phase III. New manufacturing sites, new dosage forms, and new regulatory filings all need to be reflected in the document.

In practice, quality agreements often lag behind the program. Amendments are negotiated slowly, and in the meantime the teams operate under informal understandings that have no contractual force. Building a formal review cycle into the program management plan, with responsibility assigned to a named individual on each side, is the most reliable way to prevent this.

How Ardena Approaches Quality Agreements

Ardena’s quality team produces quality agreements that are specific to each program rather than generic templates with blanks filled in. The change control classifications, notification timelines, and responsibility assignments in an Ardena quality agreement reflect the actual work being done and the regulatory context of the program.

Inside Ardena Somerset: US-Based ASD and Oral Solid Manufacturing

Why a US Site Matters

For biotech companies based in North America, time zone alignment and proximity are not trivial. A development partner eight hours ahead means decisions made in the afternoon do not get actioned until the next day. Equipment access, sample shipping, and site visits all become harder when the facility is on another continent.

Ardena’s Somerset, New Jersey site gives North American sponsors a US-based manufacturing partner with direct access to the oral solid and bioavailability enhancement capabilities the Ardena network is known for. The same scientific standards, the same quality systems, the same integrated project management, but in the same time zone.

What the Site Does

Spray Drying for Amorphous Solid Dispersions

Somerset’s core strength is spray-dried amorphous solid dispersion (ASD) manufacture. The site operates laboratory and GMP-scale spray drying equipment with closed-loop solvent recovery and secondary drying capability. The formulation team has developed ASDs across a wide range of BCS Class II molecules and polymer systems, including HPMC-AS, PVPVA, and PVP-K grades.

Development programs at Somerset start with feasibility: does this molecule respond to ASD? If yes, which polymer gives the best balance of dissolution performance, physical stability, and processability? From there, the team scales through development batches to GMP clinical manufacture within the same facility, with no formal tech transfer between scales.

Hot Melt Extrusion

HME capability at Somerset serves molecules where spray drying is less suitable, typically thermally stable APIs where a solvent-free process is preferred or where the downstream processing of a spray-dried powder presents challenges. The site’s HME expertise covers twin-screw extrusion, extrudate milling, and downstream tableting or capsule filling of the milled intermediate.

Downstream Oral Solid Manufacturing

Somerset handles the full downstream processing chain for ASD-based products: granulation where required, blending, tableting, and capsule filling. Analytical capabilities support content uniformity, dissolution testing, and stability studies from development through to GMP release.

Who Somerset Is Built For

Program TypeWhy Somerset Is a Good Fit
BCS Class II molecules needing ASD strategySpray drying and HME under one roof; formulation screening to GMP clinical batch at a single site
North American sponsors wanting US-based manufacturingSame time zone; direct site access; FDA cGMP operations; no import/export logistics for US clinical supply
Programs requiring dual US/EU clinical supplySomerset provides FDA cGMP US supply; Ghent or Pamplona provides EU GMP supply under coordinated Ardena project management
Early-phase ASD feasibility with tight timelinesRapid screening capability; formulation team responds quickly; no transatlantic coordination overhead
IND-stage oral solid programsExperience preparing CMC sections for IND submissions; regulatory team aligned with FDA expectations

The Connection to the European Network

Somerset is not an isolated US operation. It is a node in the Ardena network, with direct scientific and project management connections to Ghent, Oss, Assen, and Pamplona. For programs where solid state work needs to happen in Ghent before formulation development begins in Somerset, the handoff is internal. For programs requiring EU GMP supply for European trials alongside US supply from Somerset, a single project manager coordinates both.

That integration removes the coordination overhead that sponsors would otherwise manage themselves when using a US development house alongside a separate European manufacturer.

Inside Ardena Pamplona: High Potency, Oral Solids, and Controlled Substances

The Site Built for Difficult Molecules

Not every pharmaceutical development program involves a straightforward API in a standard formulation. Some molecules are highly cytotoxic. Some are scheduled narcotics. Some are biologically active at doses measured in micrograms. These are not unusual compounds at the edges of drug development. They are increasingly common in oncology, neurology, and rare disease pipelines.

Ardena’s Pamplona facility, which operates under the Idifarma brand within the Ardena network, is the site in the group best equipped to handle them. OEB 4 and 5 containment, controlled substance manufacturing licenses, and a formulation team experienced in the specific challenges of high-potency and low-solubility molecules make Pamplona the natural home for programs that other sites cannot take on.

HPAPI Manufacturing and Containment

Pamplona’s HPAPI infrastructure includes containment isolators for handling at OEB 4 and 5, dedicated manufacturing suites with independent HVAC systems, and validated decontamination procedures using hydrogen peroxide vapor. The site’s environmental monitoring program covers HPAPI air monitoring in work areas using validated analytical methods capable of detecting compounds at the nanogram per cubic meter level.

The containment capability supports both drug substance handling (weighing, dispensing, and transfer of HPAPI starting materials and intermediates) and drug product manufacture (blending, granulation, and tableting of HPAPI oral solid drug products). Yield accountability and mass balance documentation are maintained throughout, meeting the strict record-keeping requirements associated with potent compound manufacturing.

Controlled Substance Manufacturing

Pamplona holds manufacturing authorizations for narcotic and psychotropic substances under Spanish and European regulatory requirements. This enables the site to manufacture clinical batches of scheduled compounds including opioids, cannabinoids, and emerging psychedelic-derived therapeutics within a GMP framework that meets both EU GMP and the specific national requirements of the Spanish competent authority (AEMPS).

The site has experience managing the import and export license requirements associated with controlled substance clinical supply, and can coordinate with regulatory authorities in importing countries to ensure that cross-border shipments are compliant.

Oral Solid Formulation for Challenging Molecules

Pamplona’s formulation team specializes in molecules that present both potency and solubility challenges simultaneously. The site operates hot melt extrusion and wet milling capability for bioavailability enhancement, alongside standard granulation and tableting for more conventional formulations. High-sensitivity LC-MS/MS analytical methods are used for content uniformity and dissolution testing of HPAPI drug products where standard HPLC-UV methods lack sufficient sensitivity.

What Pamplona Is Best Suited For

Program TypeWhy Pamplona Is a Good Fit
Oncology HPAPI oral solidOEB 4-5 containment; formulation development; GMP clinical batch; high-sensitivity analytical methods
ADC payload synthesis or handlingHPAPI containment for cytotoxic warheads; controlled transfer to conjugation chemistry steps
Controlled substance developmentAEMPS manufacturing license; experience with import/export documentation; GMP batch supply for scheduled compounds
Psychedelic-assisted therapy programsSchedule 1 and 2 handling capability; experience with regulatory requirements for novel controlled substance clinical programs
BCS Class II HPAPI formulationHME and wet milling with containment; simultaneous management of solubility challenge and exposure risk
Low-dose solid dosage formsExperience with microgram-level content uniformity; validated LC-MS/MS release methods

Integration with the Ardena Network

For ADC programs, Pamplona’s HPAPI capabilities connect with the formulation and fill-finish work at Ghent and the bioanalytical program at Assen. For programs involving solid state uncertainty, Pamplona’s formulation work draws on the solid state research expertise at Ghent. For programs requiring simultaneous EU and US GMP compliance, Pamplona’s EU GMP certification and the US cGMP status of the Somerset facility together provide the geographic footprint needed for dual submissions.

Inside Ardena Assen: Bioanalysis, Clinical Supply, and Patient Kits

Where the Clinical Trial Meets the Laboratory

Ardena’s Assen facility sits at the intersection of two things that are normally separated: the bioanalytical laboratory that measures what is happening in patients and the clinical supply operation that gets the drug to them. Bringing these capabilities together under one roof is not just operationally convenient. It means the people designing the patient kit are the same people who will receive the blood samples it generates, and the lessons from one directly improve the other.

Bioanalytical Services at Assen

Small Molecule PK by LC-MS/MS

The analytical chemistry team at Assen provides fully validated LC-MS/MS methods for quantification of small molecule drugs and their metabolites in plasma, serum, urine, and other biological matrices. Studies run under GLP or GCP conditions as required, with ICH M10-compliant validation packages and ISR programs as standard.

Large Molecule and Immunogenicity Assays

The ligand-binding assay platform at Assen covers PK quantification for biologics including monoclonal antibodies, ADCs, and fusion proteins, alongside the complete tiered immunogenicity testing program for anti-drug antibody detection and characterization. MSD electrochemiluminescence and ELISA platforms are both operated, with assay development experience across a wide range of therapeutic targets.

Biomarker and Flow Cytometry

The biomarker team provides fit-for-purpose assay development and validation for pharmacodynamic, safety, and predictive biomarkers using MSD multiplex platforms, single-plex ELISA, and qPCR. The flow cytometry laboratory runs multi-parametric immunophenotyping panels for T cell subset analysis, B cell characterization, NK cell phenotyping, and MRD assessment in hematological programs.

Clinical Supply and Patient Kit Services

The clinical supply team at Assen assembles and quality-checks patient kits for clinical trials, including pre-labeled sample collection tubes, visit-specific configurations, cool pack integration, and laminated step-by-step processing instructions designed to minimize site errors. Kits are assembled to program-specific designs, with barcode labeling, personalization to patient ID where required, and IATA-compliant packaging for temperature-controlled international shipment.

Sample logistics are managed end to end: kits ship out to investigator sites, samples arrive back at Assen, chain of custody is logged from receipt, and storage conditions are verified against validated stability data before analysis proceeds. The same team that designed the kit handles the incoming samples, which means any recurring collection errors are identified and fed back to sites quickly.

The Scope of What Assen Handles

Service AreaCapabilityRegulatory Standard
Small molecule bioanalysisLC-MS/MS in plasma, serum, urine, CSF, tissueGLP / GCP; ICH M10 compliant
Large molecule bioanalysisLBA (MSD, ELISA) for mAbs, ADCs, bispecifics, fusion proteinsGLP / GCP; ICH M10 compliant
ImmunogenicityTiered ADA screening, confirmation, titration, neutralizationFDA and EMA guideline compliant
Biomarker assaysMSD multiplex, ELISA, qPCR; fit-for-purpose validationFFP approach; scope defined by intended use
Flow cytometryMulti-parametric panels; up to 15+ parameters; immunophenotyping and MRDFit-for-purpose; EuroFlow-aligned panel design
Patient kit assemblyVisit-specific kits, barcode labeling, cool pack integration, IATA packagingQuality-controlled assembly; batch records
Clinical sample logisticsReceipt, chain of custody, condition verification, storage managementGCP-compliant sample handling
Global clinical distributionIMP storage and distribution to EU and international sites; customs coordinationGDP-compliant distribution; temperature-monitored

Connecting Assen to the Rest of the Ardena Network

Bioanalytical data from Assen is interpreted alongside PK, PD, and safety data generated across the wider clinical program. For nanomedicine programs, the nanoparticle payload bioanalysis at Assen connects directly with the formulation and CQA data from Oss. For ADC programs, the ADC PK and immunogenicity assays at Assen are coordinated with the HPAPI manufacturing at Pamplona. The integration is structural, not just aspirational.

Inside Ardena Oss: The Nanomedicine Manufacturing Center

A Site Built Around Nanoparticles

Ardena’s facility in Oss, the Netherlands, is not a general pharmaceutical site that also does nanomedicines. It is a site designed from the ground up for nanoparticle development and manufacturing. The equipment, the analytical infrastructure, the technical expertise, and the quality systems are all configured around the specific demands of LNPs, polymeric nanoparticles, liposomes, and related complex nanotechnology-based products.

That focus matters. The difference between a general injectable site that can manufacture an LNP and a dedicated nanomedicine site is the same as the difference between a GP and a specialist. The specialist has seen the edge cases, solved the unexpected problems, and built the institutional knowledge that makes the process more reliable.

What Oss Offers

LNP Development and GMP Manufacturing

The formulation team at Oss develops LNP products from initial lipid screening through to GMP clinical batch manufacture. Microfluidics and nanoparticle extrusion platforms are available at development and GMP scale, with integrated tangential flow filtration (TFF) capability for buffer exchange and concentration. The site has experience with mRNA, siRNA, and small molecule LNP products across a range of ionizable lipid systems.

Polymeric and Other Nanoparticle Systems

Beyond LNPs, the Oss team has formulation capabilities for PLGA and other biodegradable polymer nanoparticles, nanosuspensions prepared by wet milling, and lipid-polymer hybrid systems. The common thread is that the site is equipped for the characterization-intensive development work that nanoparticle products require, with DLS, NTA, RiboGreen encapsulation efficiency assays, and in vitro release testing all available in-house.

Analytical Characterization

The analytical team at Oss provides the full suite of physicochemical characterization required for LNP and nanoparticle drug product development, including particle size and PDI by DLS, zeta potential, encapsulation efficiency by RiboGreen or equivalent, lipid composition by HPLC, and mRNA integrity by gel electrophoresis. The team develops the analytical methods needed for GMP batch release and regulatory filing alongside the formulation program.

The Programs Oss Is Best Suited to Support

Program TypeWhy Oss Is a Good Fit
mRNA LNP therapeuticsDedicated microfluidics and TFF equipment; mRNA integrity and potency testing; GMP clinical batch capability
siRNA or ASO LNPNucleic acid encapsulation experience; encapsulation efficiency optimization; GMP manufacturing at clinical scale
Polymeric nanoparticle sustained releasePLGA and polymer nanoparticle development; in vitro release testing; GMP scale-up
Nanosuspension for BCS Class IIWet milling with particle size characterization; downstream processing to solid dosage form
LNP regulatory filing supportIntegrated regulatory team; experience with Module 3 CMC for LNP products; familiarity with FDA and EMA expectations
Scale-up from development to GMPSame facility and team at both scales; no formal tech transfer; process parameters maintained consistently

Connection to the Broader Ardena Network

Oss does not work in isolation. For ADC programs where the payload requires HPAPI containment, the formulation work at Oss connects with the Pamplona facility’s containment manufacturing capability. For bioanalysis of nanoparticle payloads, Oss works alongside the bioanalytical team at Assen. For aseptic fill-finish of finished nanoparticle drug products, Oss programs can connect to the sterile manufacturing infrastructure at Ghent.

Inside Ardena Ghent: Oral Solids, Sterile Manufacturing, and Solid State Research

The Site That Does the Most

Of all Ardena’s European sites, Ghent carries the broadest scope. It is where solid state research happens. It is where oral solid development and manufacturing runs. It is also where sterile injectables, lyophilized products, and complex parenteral formulations are developed and manufactured under GMP. For a program that spans multiple dosage forms or that needs solid state characterization feeding directly into drug product development, Ghent is where those conversations happen in one building.

Solid State Research

The solid state research group in Ghent is dedicated to pre-formulation work: polymorph screening, salt and co-crystal screening, hydrate and solvate assessment, and the characterization of physical and chemical properties that determine how a molecule will behave in a formulation. The team works with XRPD, DSC, TGA, DVS, and Raman spectroscopy as its core analytical toolkit.

What makes the Ghent setup particularly effective is the direct connection between solid state research and drug product development. The scientist who identifies the optimal salt form is part of the same organization as the scientist who will formulate the tablet. Decisions made in pre-formulation are informed by what the downstream process can accommodate, rather than handed over in a report and interpreted by someone else months later.

Oral Solid Development and Manufacturing

Ghent’s oral solid development team covers the full formulation development pathway for tablets and capsules: pre-formulation, formulation screening, process development, and scale-up to GMP clinical batch manufacture. The site operates batch and continuous manufacturing capability for tableting, granulation (wet and dry), film coating, and capsule filling.

PAT tools including near-infrared spectroscopy are integrated into the manufacturing environment, supporting blend uniformity monitoring and granulation endpoint determination. The team has experience with immediate release, modified release, and enteric-coated dosage forms, and with challenging APIs including poorly soluble compounds that require bioavailability enhancement strategies.

Sterile Manufacturing and Lyophilization

The sterile manufacturing suite at Ghent provides aseptic fill-finish for vials and ampoules under Grade A conditions within a RABS environment. Lyophilization capacity supports both conventional small molecule injectables and biologics requiring freeze-drying for stability.

The sterile manufacturing team has experience with complex injectable formulations including liposomal products and nanoparticle drug products requiring the specific process controls and in-process monitoring discussed in the broader nanomedicine fill-finish program. Environmental monitoring and media fill qualification are maintained to EU GMP Annex 1 standards.

Analytical and Regulatory Support

Ghent’s analytical teams provide method development and validation for both oral solid and injectable products. The regulatory team in Ghent contributes to CMC dossier preparation for IND and IMPD submissions, with particular expertise in the solid state and oral solid sections of Module 3.

What Ghent Is Particularly Well Suited For

Program TypeWhy Ghent Is a Good Fit
Early-phase oral solid with solid state uncertaintySolid state research and formulation development in the same site; findings translate directly into formulation decisions
Poorly soluble oral APIHME, wet milling, and lipid formulation capabilities alongside solid state analytics
Sterile injectable for Phase IAseptic fill-finish under EU GMP; lyophilization available; experienced with small clinical batches
Liposomal or nanoparticle injectableSpecialist experience in nanoparticle fill-finish process development; particle size monitoring during filling
Program needing dual IMPD/IND filingEU GMP certified site; regulatory team experienced in both jurisdictions
Continuous manufacturing evaluationBatch and continuous manufacturing capability; PAT tools in place; team experienced in Q13 requirements

Working with the Ghent Team

Projects based primarily at Ghent are managed by dedicated project managers who coordinate work across the solid state, formulation, analytical, and manufacturing teams within the site. For multi-site programs, the Ghent project manager integrates with counterparts at Oss, Assen, Pamplona, and Somerset to ensure that the work is coordinated and that data flows between teams without friction.

Navigating the European CDMO Landscape: EU vs. US Regulations

Why the Geography of Your CDMO Partner Matters

The choice to develop a drug in Europe rather than the United States is not just a matter of geography. It affects your regulatory strategy, your manufacturing standards, the reference points in your CMC dossier, and ultimately the timeline to your first clinical data. For many biotech companies, particularly those seeking approval in both markets, understanding the interplay between EMA and FDA requirements from the outset saves significant rework later.

Key Regulatory Differences: EMA vs. FDA

Regulatory ElementEMA (European)FDA (United States)
Pre-clinical to Phase I submissionIMPD (Investigational Medicinal Product Dossier) in CTD formatIND (Investigational New Drug application)
GMP standardEU GMP (EudraLex Vol 4)cGMP (21 CFR Parts 210/211)
Regulatory inspection frameworkNational Competent Authorities + EMAFDA CDER/CBER inspection
CMC data expectations at Phase IPragmatic; fit-for-purpose with defined limitationsSimilar; FDA responsive to first-cycle CMC deficiency letters
Pediatric obligationPaediatric Investigation Plan (PIP) requiredPediatric Study Plan (PSP) under PREA
Advanced therapy productsATMP Regulation (1394/2007), CAT committeeBLA/NDA process through CBER
Clinical trial authorizationSingle CTA per Member State or EU CTA (CTIS)IND filing covers US clinical sites

The Advantages of Developing in Europe

EU GMP Is Recognized Globally

EU GMP certification is accepted by a wide range of health authorities beyond Europe, including those of Australia, Canada, Japan, and many emerging markets. Developing under EU GMP from the outset means your manufacturing data is audit-ready for more markets without additional inspection overhead. The EU GMP mutual recognition agreements with the FDA and other agencies further reduce duplication for globally-focused programs.

Parallel IMPD/IND Submissions Are Achievable

For companies targeting both EU and US clinical trials, a well-structured CMC dossier developed in Europe can form the basis of both an IMPD and an IND submission. The CTD format (Common Technical Document) was designed precisely to enable this, with Module 3 quality data common to both applications. A CDMO with experience in dual submissions can build that efficiency into the development plan from the start.

Cost-Effective Early Development

For pre-clinical and Phase I work, European CDMOs often offer highly competitive pricing relative to comparable US facilities, with equivalent or superior scientific capabilities. For a capital-efficient biotech, that cost advantage at the development stage can meaningfully extend the runway to the next funding milestone.

How Ardena Supports Multi-Jurisdictional Programs

Ardena’s facilities are authorized under EU GMP and operate within regulatory environments that are routinely inspected by national competent authorities across Belgium, the Netherlands, Spain, and the US FDA. The organization has experience preparing CMC sections for both IMPD and IND submissions, and can advise on where regulatory strategies diverge between jurisdictions.

For programs seeking to run EU and US trials in parallel, Ardena’s regulatory team can help structure the CMC development plan to produce data that satisfies both agencies without duplicating the experimental work